LATE ONSET FABRY’S DISEASE MIMICKING HYPERTROPHIC CARDIOMYOPATHY
European Heart Journal Supplements
Abstract
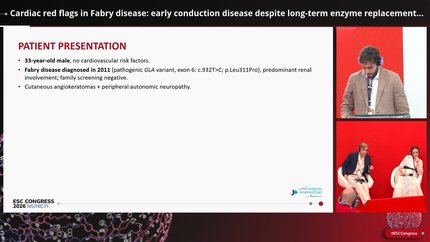
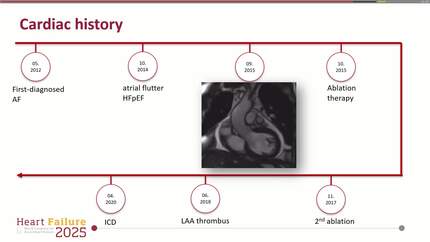
Fabry disease (FD) is rare genetic syndrome which determines a lysosomal storage disorder caused by pathogenic variants in the alpha–galactosidase A (GLA) gene that lead to alpha–galactosidase A (AGAL– A) enzyme activity disfunction.The fenotipic effect is a left ventricular hypertrophy (LVH) that is frequently misdiagnosed as hypertrophic cardiomyopathy (HCM).This disease in still unfamiliar for the most part of cardiologists cause of the fenotipic familiarities with other more common and wellknown diseases and according to recent data the prevalence of FD among unexplained LVH is 0.93%.These characteristics drive to a heavy under diagnosis of a disease with many therapy chances. The prompt initiation of these treatments, subsequently an early diagnosis, is actually considered the best strategy to improve the natural course of the disease.The critical evaluation of the clinical history and basic cardiomiopaty assessments could drive the cardiologist to the diagnosis.Precisely, we present an example of three family pedigree discovered in a couple of months subsequently the unmasking of proband male patients with decennary misdiagnosis of HCM. The ambulatory suspect of non sarcomeric disease in LVH drove us to critically reexaminate EKGs and previous CMR scans finding respectively PR shortage associated with right bundle block and basal inferolateral late gadolinium enhancement (figure 1). In a proband it is not possible to perform MRI with gadolinium due to the presence of renal failure and the diagnostic suspicion arose by analyzing the T1 mapping signal (figure 2).The score ID FABRY–HCM were applied resulting suggestive too.Having thus corroborated the suspicion, blood spot tests were performed finally detecting a very low AGAL–A activity. The following GLA genetic tests identified the tipical mutation of late onset Fabry’s disease (p.N215Ser). Fulfilling the consensus criteria for the therapy initiation, the patients were treated with enzyme replacement therapy. We completed families predigree identifying other six famale gene mutation carriers with almost preserved AGAL–A activity. The diagnosis of FD allowed us to perform the specific therapy in affected patients hopefully effecting their quality of life and their prognosis and to bring out healthy carriers.The initial suspect questioning a previous diagnosis starting to simple clinical and routine strumental elements were crucial.
Contributors
L Rivetti
Author
F Bullian
Author
R Mantovan
Author
G Allocca
Author
M Centa
Author
F Bagato
Author
N Sitta
Author
M Gallucci
Author
G Carraro
Author
N Vitturi
Author
L De Michieli
Author
You may be interested in